PSUR vs PMSR: Understanding Post-Market Surveillance Reporting Under EU MDR
Summary Under EU MDR 2017/745, post-market surveillance is no longer a passive requirement. It is [...]

Under EU MDR 2017/745, post-market surveillance is no longer a passive requirement. It is an active, continuous process of collecting, analysing, and evaluating device data after the product reaches the market because pre-market testing cannot identify every rare flaw or real-world risk. Effective postmarket surveillance helps manufacturers prevent harm, meet legal requirements, maintain market access, and reinforce buyer trust. In practice, many manufacturers underestimate how closely post-market documentation is scrutinised during Notified Body reviews.
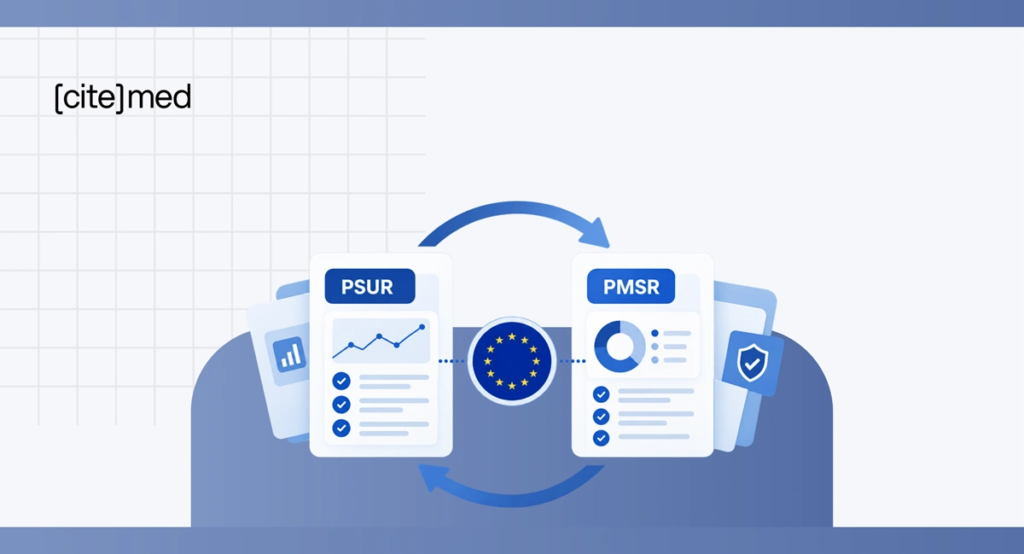
Two key reporting tools within the MDR post-market surveillance system are the Periodic Safety Update Report (PSUR) and the Post-Market Surveillance Report (PMSR). While both summarise post-market safety information, they apply to different device risk classes and involve different levels of analysis and regulatory oversight, with PSURs used for Class IIa, IIb, and III devices and PMSRs for Class I devices. For regulatory affairs, quality assurance, clinical, and medical writing teams at medical device, diagnostics, combination product, and digital therapeutics companies—especially startups, small-to-mid sized manufacturers, CROs, and independent consultants—understanding that distinction is essential to producing audit-ready documentation. This guide explains the PSUR vs PMSR difference, outlines EU MDR reporting requirements, shows where manufacturers commonly make mistakes, and connects postmarket surveillance with clinical evaluation and vigilance so your reporting system is inspection-ready.
The Periodic Safety Update Report (PSUR) is a comprehensive post-market safety evaluation required for Class IIa, IIb, and Class III medical devices under EU MDR Article 86. It includes detailed benefit-risk analysis, clinical data review, and is subject to Notified Body oversight.
The Post-Market Surveillance Report (PMSR) is a higher-level summary document required for Class I devices under EU MDR Article 85. It provides an overview of post-market surveillance findings and is primarily maintained as internal documentation within the manufacturer’s quality management system, since PMS is an essential component of the quality management system for medical devices and helps medical device manufacturers document ongoing safety monitoring.
The critical distinction is not just the document format, but that PMS provisions should be proportionate to the risk class of the device, which is why PSUR depth exceeds PMSR depth. It is the level of analytical depth, the regulatory oversight involved, and the frequency of update required. Using a PMSR where a PSUR is required is a compliance failure. Producing a PSUR that reads like a PMSR, listing data without critically evaluating it, is one of the most common reasons post-market submissions are challenged during Notified Body review.
Under EU MDR, post-market surveillance refers to the systematic collection, monitoring, and analysis of data on a medical device after it has been placed on the market. The goal is to ensure that devices continue to perform as intended and remain safe throughout their lifecycle. Under MDR, PMSRs apply to Class I devices, while PSURs are required for higher-risk devices under Articles 85 and 86.
Manufacturers are expected to implement structured PMS systems capable of identifying safety signals, detecting emerging risks, and helping maintain the benefit-risk ratio through dynamic risk management. These systems rely on multiple data sources including complaints, user feedback, adverse events reports, clinical literature, and post-market clinical follow-up studies.
Key activities within the PMS framework typically include complaint monitoring and trend analysis, vigilance reporting of serious incidents, post-market clinical follow-up studies, literature surveillance and clinical data review, and evaluation of real-world device performance.
The results of these surveillance activities must be systematically documented and periodically summarised through formal reports. Depending on device classification, manufacturers must prepare either a PMSR or a PSUR and report relevant safety and performance data to regulatory authorities, reflecting broader global requirements for manufacturers to monitor products after launch.
The Post-Market Surveillance Report is a summary document required for Class I medical devices. Its primary purpose is to provide a structured overview of post-market surveillance findings and confirm that the device continues to meet applicable safety and performance requirements.
Because Class I devices are considered lower-risk products, the PMSR is generally less detailed than higher-risk safety reports. It summarises the results of PMS activities including tracking adverse events, collecting customer complaints, monitoring product performance, complaint trends, vigilance data, and any relevant findings from literature monitoring or post-market evaluations.
Although the PMSR is typically maintained as internal documentation within the manufacturer’s quality management system, it must still be kept up to date and available for regulatory inspection, with strong internal communication channels for logging complaints and feedback in a timely manner, as competent authorities and Notified Bodies may review these reports during audits or regulatory inspections.
The report should demonstrate that manufacturers are actively monitoring device performance and that no new safety concerns have emerged that would alter the device’s risk benefit profile.
The Periodic Safety Update Report is a more comprehensive post-market surveillance document required for higher-risk devices. Under EU MDR Article 86, PSURs are a mandatory requirement for Class IIa, IIb, and III devices. In practice, one of the most common reasons PSURs fail Notified Body review is the absence of a clear, structured benefit-risk evaluation, with data presented descriptively rather than critically analysed. PSUR-ready systems should operate under a formal pms plan that defines objectives, data sources, analysis frequency, reporting format, and how outputs feed into technical documentation.
Unlike the PMSR, the PSUR provides a detailed analysis of post-market safety and performance data. It evaluates the device’s risk benefit balance using information collected through PMS activities, including complaint trends, adverse event reports, PMCF studies, and other relevant clinical data sources, and this real-world evidence also supports continuous product improvement.
PSURs must also include information on sales volumes, population exposure estimates, and usage frequency, as these factors help regulators understand the scale of device use and contextualise safety findings. The analysis should also support timely corrective action when safety risks are identified, including CAPA where appropriate. Regulators need context, not just raw incident data, to assess real-world risk.
Reporting frequency depends on device classification. Class IIa devices require a PSUR updated at least every two years. Class IIb and Class III devices require annual updates. For many higher-risk devices, PSURs are reviewed by Notified Bodies, making them an important component of regulatory oversight and lifecycle safety evaluation.
Despite clear MDR guidance, manufacturers sometimes misunderstand the differences between PMSR and PSUR requirements.
One common misconception is that a PMSR can be used for devices that require a PSUR. Higher-risk devices must comply with the more stringent PSUR reporting requirements. This is not a grey area under EU MDR.
Another frequent issue involves misunderstanding the reporting frequency associated with PSURs. Because these reports must be updated regularly, manufacturers must establish internal systems capable of continuously collecting and analysing safety data. Treating PSUR preparation as a reactive exercise rather than an ongoing process produces reports that are out of date at the point of submission.
Manufacturers may also underestimate the level of analysis expected in PSUR documentation. Regulators expect PSURs to include meaningful safety evaluations, including signal detection through analysis of trends and unexpected safety signals, rather than simply listing surveillance data. If there is one pattern that consistently emerges during regulatory review, it is that manufacturers underestimate the level of interpretation expected, not just data reporting. A PSUR that presents complaint numbers without contextualising them against population exposure, or that reports PMCF findings without connecting them to the benefit-risk conclusion in the CER, and without integrating those findings with the risk management file rather than reviewing them in isolation, will generate Notified Body queries regardless of how complete the underlying data appears. This analysis creates a feedback loop for product development and, where needed, changes to the device’s design.
For a full breakdown of how documentation gaps affect regulatory submissions see our article on reasons Clinical Evaluation Reports are rejected.
For manufacturers operating under MDR, understanding the distinction between PMSR and PSUR is essential for effective regulatory strategy. These reports are not merely administrative documents; they represent critical tools for demonstrating ongoing device safety and performance.
Establishing a robust PMS system is key to generating the data needed for both reports, but this depends on effectively integrating complaint handling systems, vigilance processes, and clinical evaluation activities, including PMCF as a proactive methodology using studies designed to confirm product safety, into a comprehensive lifecycle monitoring strategy.
Companies should also plan for the analytical workload associated with PSUR preparation, particularly for higher-risk devices. This typically involves establishing structured processes through a formal plan that defines objectives, data sources, analysis frequency, and reporting format, with provisions proportionate to the device risk class, for ongoing data collection, signal detection, and periodic data review. Effective documentation systems can significantly reduce the complexity of preparing these reports and ensure that safety information is readily available when required.
Notified Bodies reviewing PSURs focus specifically on three things:
A clear and structured benefit risk assessment, not a list of incidents but a genuine assessment of whether the device’s clinical benefits continue to outweigh its risks given the totality of available post-market evidence. This evaluation must reference the current state of the art and be consistent with the conclusions presented in the CER.
Justification of observed trends and their clinical relevance. If complaint rates have increased, the PSUR must explain why, assess whether this represents a genuine safety signal, and document what action was taken or why no action was required.
Clear linkage between PMS data and clinical evaluations. The PSUR should connect directly to the CER, the risk management file, and PMCF documentation in a way that tells a coherent safety story across the full technical file.
| Aspect | PSUR | PMSR |
|---|---|---|
| Device class | Class IIa, IIb, III | Class I |
| Regulatory basis | EU MDR Article 86 | EU MDR Article 85 |
| Purpose | Detailed safety evaluation | PMS summary |
| Frequency | Annual (IIb/III) or every two years (IIa) | Updated as needed |
| Regulatory review | Reviewed by Notified Bodies; psur submission also becomes available via EUDAMED from 2026 | Primarily internal |
| Level of analysis | Comprehensive benefit-risk evaluation | High-level summary |
| Sales and exposure data | Required | Not required |
| PMCF integration | Required | Recommended |
| CER linkage | Explicit and documented | General alignment |
Note: EUDAMED registration is mandatory by May 28, 2026.
Specific to PSUR:
Specific to PMSR:
Post-Market Surveillance (PMS)
The systematic process of collecting, analysing, and evaluating data on a medical device after market entry. Under EU MDR, PMS is a continuous active obligation rather than a periodic reporting exercise.
Periodic Safety Update Report (PSUR)
A comprehensive post-market safety document required for Class IIa, IIb, and Class III devices under EU MDR Article 86. It includes detailed benefit-risk evaluation, clinical data analysis, and exposure data, and is subject to Notified Body review.
Post-Market Surveillance Report (PMSR)
A summary post-market document required for Class I devices under EU MDR Article 85. It provides a high-level overview of surveillance activities and is primarily maintained as internal documentation within the QMS.
Benefit-risk evaluation
The formal assessment of whether the clinical benefits of a device outweigh its risks in the context of the available post-market evidence. In a PSUR, this evaluation must be analytically supported and consistent with the conclusions presented in the CER.
Post-Market Clinical Follow-Up (PMCF)
The structured process through which manufacturers actively collect clinical data on their device after market entry. PMCF findings feed directly into both the PSUR and the CER and are a required component of the PMS framework for higher-risk devices.
Vigilance reporting
The process of reporting serious incidents and field safety corrective actions to competent authorities within defined timelines under EU MDR. Vigilance data forms a key input to both PSUR and PMSR documentation.
Signal detection
The process of identifying emerging safety concerns from post-market data before they become reportable events. Effective signal detection requires structured analysis of complaint trends, vigilance data, literature findings, and PMCF outcomes over time.
The most consistent pattern in PSUR failures during Notified Body review is not missing data. It is missing interpretation. Manufacturers collect complaint rates, vigilance reports, and PMCF findings and present them accurately, but do not take the next step of evaluating what those findings mean for the device’s ongoing benefit-risk profile. A PSUR that reports a 12% increase in device-related complaints without explaining whether this represents a genuine safety signal, a change in reporting behaviour, or an expected consequence of increased market penetration will generate a deficiency letter regardless of how thoroughly the data itself is documented. The analytical layer is not optional. It is the point of the document.
The PSUR (Periodic Safety Update Report) is required for Class IIa, IIb, and Class III devices under EU MDR Article 86 and provides a comprehensive safety evaluation including benefit-risk analysis, clinical data review, and exposure data. It is subject to Notified Body review. The PMSR (Post-Market Surveillance Report) is required for Class I devices under Article 85 and provides a higher-level summary of post-market surveillance activities. It is primarily maintained as internal documentation. The critical difference is the depth of analysis required and the level of regulatory oversight involved.
Under EU MDR Article 86, Class IIa devices require a PSUR updated at least every two years. Class IIb and Class III devices require annual updates. These timelines are minimum requirements. Where significant new safety data emerges, the PSUR should be reassessed regardless of the scheduled update cycle. For more detail on building a structured post-market surveillance system that supports ongoing PSUR preparation, see our dedicated article on this topic.
The most common reason is the absence of a structured benefit-risk evaluation. Many PSURs present surveillance data accurately but fail to critically analyse what that data means for the device’s ongoing safety profile. Other frequent issues include weak linkage between PSUR conclusions and the Clinical Evaluation Report, inadequate contextualisation of complaint trends against population exposure, and PMCF findings that are reported but not integrated into the benefit-risk conclusion.
A PSUR draws on the full range of post-market surveillance activities including complaint monitoring and trend analysis, vigilance reporting data, PMCF study findings, literature surveillance results, real-world performance data, sales volume and population exposure estimates, and registry data where available. The quality of the PSUR depends directly on the quality and completeness of the underlying PMS system that generates this data.
No. The PMSR is primarily maintained as internal documentation within the manufacturer’s quality management system. However, it must be kept current and made available for inspection by competent authorities or Notified Bodies if requested during audits or regulatory reviews. Manufacturers should not treat the PMSR as a document that will never be scrutinised.
Post-market surveillance findings must feed directly into the Clinical Evaluation Report. The CER must be updated to reflect new evidence generated through PMS, PMCF, and vigilance activities. Where PMS data and CER conclusions are developed in isolation, inconsistencies emerge that Notified Bodies identify during conformity assessment. For a full breakdown, see our article on reasons Clinical Evaluation Reports are rejected.
If you are preparing a PSUR or PMSR, reviewing your post-market surveillance system, or building a more structured approach to lifecycle evidence management under EU MDR, Citemeds can help. Get in touch to discuss your requirements.
Literature reviews, CERs, post-market surveillance — in one platform.